Cell-cultivated products: toxicology and growth media assessment
Published 30 September 2026
Applies to England, Scotland and Wales

© Crown copyright 2026
This publication is licensed under the terms of the Open Government Licence v3.0 except where otherwise stated. To view this licence, visit nationalarchives.gov.uk/doc/open-government-licence/version/3 or write to the Information Policy Team, The National Archives, Kew, London TW9 4DU, or email: psi@nationalarchives.gov.uk.
Where we have identified any third party copyright information you will need to obtain permission from the copyright holders concerned.
This publication is available at https://www.gov.uk/government/publications/cell-cultivated-products-toxicology-and-growth-media-assessment/cell-cultivated-products-toxicology-and-growth-media-assessment
1. Scope of this guidance
This supplemental guidance applies specifically to cell-cultivated products regulated as novel foods in Great Britain (GB).
Cell-cultivated products cover a variety of foods that can be made using a production process without slaughter or traditional farming and agricultural practices. Cells isolated from animals or plants, including cells from meat, seafood, fat and offal, or eggs, are grown in a controlled environment, and then harvested to make a final food product.
This supplemental guidance is inclusive of products derived from both invertebrate and vertebrate animal cells such as:
- mammalian
- fish
- shellfish
- avian cells, particularly those intended to replicate the appearance and characteristics of conventional meat or seafood
You should comply with Regulation (EU) 2015/2283 when applying for market authorisation for cell-cultivated products. This requires evidence to identify and characterise hazards and demonstrate effective mitigations are in place to ensure consumer safety.
This supplemental guidance addresses hazards associated with growth media components and provides considerations for the toxicological assessment of the cell-cultivated products as novel foods.
The guidance is not intended to prescribe a single approach or provide an exhaustive account of all methodologies. As scientific understanding and risk assessment approaches continue to advance, alternative approaches may be appropriate, provided they are:
- scientifically justified
- generate reliable and supportive data
- comply with the applicable novel food regulatory requirements
2. General considerations for your safety assessment
You should support all safety assessments with appropriate scientific evidence. Analyses should be conducted by competent facilities using nationally or internationally recognised and appropriate accredited methods.
Your application should include a full description of the methods employed. Test materials used in any studies should be representative of the product intended to be marketed. You should also provide copies of the certificates of analysis.
You should follow, and provide evidence of using, official guidelines and quality systems such as Organisation for Economic Co-operation and Development (OECD) test guidelines or Good Laboratory Practise (GLP).
Testing should use accredited methods where available. Where not available, testing should be done in an accredited facility fit for the purpose of the methods.
Where neither an accredited method nor facility are available, you may consider ‘in-house’ methods. These should be fully described standardised internal methods with appropriate controls. You should justify why you’ve used these methods in your application.
You should use this guidance alongside:
- assimilated Regulation (EU) 2015/2283
- the 2016 technical guidance from EFSA for novel food applications
- other cell-cultivated product specific supplementary guidance published by the Food Standards Agency (FSA) and Food Standards Scotland (FSS)
The FSA and FSS review all applications to make sure they meet regulatory standards. We may ask for more information during the approval process if we identify any gaps or uncertainties in the application.
3. Assessing growth media
In your novel food application, you must assess the safety of growth media components used to produce cell-cultivated products. This section explains the scientific and regulatory requirements you should use.
The guidance supplements assimilated Regulation (EU) 2015/2283 and is intended to be used in conjunction with the 2016 EFSA guidance for novel foods.
Some growth media substances are commonly found in conventional foods. Others may have no prior history of consumption.
To support the safety assessment of these components, you should use a structured and proportionate framework. For example, prioritise your assessment based on each component’s risk level (using existing safety data).
You should consider the potential for growth media components to bioaccumulate within cells and assess whether this could contribute to the toxicological profile of the final novel product.
3.1 Listing your growth media components
You should provide a structured, complete list of all growth media components used at every stage of production. This includes:
- cell isolation
- proliferation
- differentiation
- harvest
- any post-harvest operations where exposure to media components could occur
This includes, but is not limited to:
- basal media and culture formulations
- supplements, such as amino acids, vitamins, lipids, minerals, trace elements
- growth factors, cytokines, hormones (including recombinant proteins) and small molecules
- antimicrobials and antifungals
- processing aids, surfactants, pH regulators, anti‑foam agents, chelators
- cryopreservation media and thawing solutions
If proprietary ingredients are supplied by third-party manufacturers, you should provide information about the supplier. Make sure the supplier submits comprehensive component safety information directly to the FSA, in confidence, to support the risk assessment.
3.2 Documenting your component inventory
For each component used at any stage of the production process, you should provide a complete and structured set of information sufficient to support traceability and safety assessment.
You should include the following information for every component in the register:
- stages of production at which the component is used
- substance identity, including chemical name and chemical abstracts service (CAS) number
- technological properties in the production process and, where relevant, in the final novel food
- quality and quantity of component intended use, for example, food, pharmaceutical or medical, including a justification when a non-food‑safe component is used
- specification, certificates of analysis, technical data sheets and purity profile, including impurity limits and microbiological criteria
- if varied growth media or solutions are used at different stages throughout the process, indicate their composition (including stock concentration and working concentration) at each stage
- if relevant, quantification of the levels of the substance present in the final novel product, supported by standardised analytical data
- maximum safe residual concentration justified with toxicological evidence, applicable health-based guidance values, scientific literature, and validated safety information
- an assessment of consumer exposure to residues of growth media components in the final novel product, in relation to relevant health-based guidance values
- analytical method details, including method type, matrix validation, Limit of Detection, Limit of Quantitation and any other uncertainty information
- history of safe use and supporting references, including regulatory evaluations or published scientific evidence, where available
This list is not exhaustive. You can provide further information if it is relevant to support the safety assessment.
All analytical determinations should be performed using validated, sensitive methods appropriate for the cell-cultivated product matrix.
Where a substance is not detected, you should report the Limit of Detection and Limit of Quantitation to demonstrate confidence in the non-detection outcome.
3.3 Growth media components produced through recombinant DNA technology
In some cases, cell lines may be immortalised through genetic modification. For more information on genetically modified cells and the appropriate regulatory pathway see the identity, production and microbiology supplementary guidance published by the FSA and FSS.
Where media components, growth factors or enzymes are produced using recombinant DNA technology, you should provide additional molecular and safety information. This should be enough information to assess potential risks associated with the expression system and resulting product.
Your application should include:
- scientific name of the host organism, for example a microorganism, plant, or other expression system
- a detailed description of the genetic modification process, including all inserted, deleted, or modified genetic elements and their functional roles
- for recombinant plants, an assessment of host plant safety, including any known toxicity or allergenicity
- for recombinant microorganisms, an assessment of organism safety and pathogenicity, demonstrating absence of virulence factors or toxin-producing pathways, absence or non-transferability of antimicrobial resistance genes and evaluation of potential for horizontal gene transfer
- confirmation of whether the host organism has a history of safe use for producing recombinant substances used in foods
- for recombinant proteins, the quantity used within production, primary sequence data, characterisation demonstrating equivalence (or near equivalence) to the native protein, and assessment of any added purification tags or sequence modifications, with evaluation of allergenicity and toxicity
- evaluation of whether the recombinant component may give rise to any additional food safety hazards
- for recombinantly produced components, evidence demonstrating the absence of viable production organisms and residual recombinant DNA in downstream materials, where relevant to the final novel product
3.4 Specific categories and requirements
3.4.1 Nutrients and salts
For amino acids, vitamins, minerals and trace elements, you should declare:
- the grade
- specification data
- impurity limits
- expected maximum residual concentrations
You should show that any residues in the final food are safe and do not have a negative nutritional impact.
3.4.2 Cell signalling molecules, biologics and small molecules
You should provide comprehensive characterisation and safety evidence for all bioactive signalling molecules and small molecules used in the production of the novel food. This might include growth factors, cytokines and hormones.
This includes both naturally derived and recombinant forms.
You should provide:
- identification of each bioactive substance, including its molecular identity, species of origin, production method (including the host system where recombinant), and relevant structural features such as post-translational modifications
- a description of the manufacturing and processing steps that contribute to the removal, degradation or inactivation of these substances
- data on residual levels in the final novel food, generated using appropriately sensitive and specific analytical methods, or Limit of Detection and Limit of Quantitation where absence is demonstrated
- an assessment of the potential biological activity of any residual material following processing and consumption relevant conditions (for example, heat treatment or digestion)
- evidence demonstrating that the component presents no detectable biological active in the novel food and representative final products as intended to be consumed
3.4.3 Veterinary medicines
You should provide verifiable evidence demonstrating that source animals have complied with statutory veterinary medicines withdrawal periods. Evidence may include treatment records, veterinary certification, and residue testing data where appropriate.
See our allergenicity and nutrition guidance for more information.
3.4.4 Processing aids
You should provide complete characterisation and safety justification for all processing aids used during production.
Processing aids include, but are not limited to:
- surfactants
- chelators
- solubilising agents
- anti‑foam agents
- filtration enhancers
- pH modifiers
- any auxiliary substances used to facilitate manufacturing steps
To support regulatory assessment, you should:
- provide the chemical identity and CAS-number, along with the quality grade, for example, food-safe, pharmaceutical grade (where a non–food-grade material is used, provide a robust safety justification)
- submit full specification data for all processing aids used at any stage of production
- provide validated analytical data quantifying residual levels in the final novel product, or supply a scientifically justified removal rationale supported by process controls (for example, washing, filtration, or centrifugation)
- demonstrate that the substance is effectively removed or reduced to safe levels, with justification based on health-based guidance values
For more information on process aids and additives, see our cell-cultivated product regulatory authorisation guidance.
Scaffolds and microcarriers
You should provide comprehensive characterisation and safety assessment for all scaffolds, microcarriers or structural materials used during any stage of the production process.
These materials may serve as a surface for the cellular attachment to support growth. They can influence product integrity, residual composition or safety and therefore require comprehensive regulatory scrutiny.
Edible scaffolds and microcarriers
Where the scaffold or microcarrier is intended to remain in the final product, you should provide:
- full composition and material identity
- quality grade (for example, food-safe) with justification where non-food‑safe materials are used
- specifications, including physical and chemical properties, and relevant microbiological standards
- quantification of the residue level of the scaffold or microcarrier present in the final novel food, supported by validated analytical data
- a narrative demonstrating that its presence does not introduce nutritional, chemical, physical or toxicological risks
Non‑edible scaffolds and microcarriers
Where the scaffold or microcarrier is not intended to remain, you should provide:
- evidence of complete removal
- details of the process and the process controls (for example, filtration, washing, dissociation, demonstrating reliable removal)
Where you cannot demonstrate complete removal, you should provide validated residue testing of the scaffold and its components.
You should assess the potential for scaffolds and microcarriers to release:
- leachables, for example, plasticisers
- extractables under processing or digestion conditions
- degradation or breakdown products released through shear, enzymatic activity, pH shifts, or thermal or chemical processes
- particulate matter, including microplastics, generated through processing
For more information, including where additional authorisations are likely to be needed, see our cell-cultivated product regulatory authorisation guidance.
3.4.6 Water quality
You should provide clear evidence that the water used throughout the production process meets safety and quality standards appropriate for food production in the UK.
This includes water used for media preparation, cell washing, cleaning, process operations, and as an ingredient.
You should demonstrate control of chemical contaminants by residue testing in the final novel product. You may refer to assimilated Regulation No 1881/2006 for indicative lists of contaminants and associated maximum levels.
Where relevant, you should also consider recent updates, for example, the 2023 amendments under EU law.
For more information on potable water requirements and alternative washing substances, see our cell-cultivated products hygiene guidance.
3.4.7 Food contact materials and equipment
Food contact materials (FCMs) are materials and articles that come into contact with food during its production, processing, storage, preparation or serving.
You should provide a clear description and safety assessment of all FCMs and equipment that contact the product, culture media or intermediates during any stage of cell-cultivated product production. This includes:
- cell isolation
- expansion
- differentiation
- harvesting
- downstream processing
- final novel product handling
The EFSA guidance on FCMs (2017) contains further guidance.
You should provide an inventory of all materials and equipment that may contact cells, media, scaffolds, or final novel products.
For each material, you should provide:
- material identity and composition (for example, polyethylene, silicone or stainless steel)
- certification or declaration of compliance with applicable FCM regulations or standards
- intended function and stage of use in the production process
3.4.8 Residuals strategy and analytical expectations
When considering the analytical strategy for all growth media components, you should include a clear, evidence-based justification in their dossier for identifying and quantifying any residual substances that carry over into the novel food.
Pay attention to components with a higher likelihood of persistence. This includes substances that are lipophilic, chemically stable, or biologically active. You should:
- identify all inputs with plausible carryover into the final novel food, and target those with toxicological relevance, including cleaning agents or other processing materials with potential incidental food contact
- use validated analytical methods appropriate for cell-cultivated product matrices, for example, liquid chromatography with tandem mass spectrometry (LC–MS/MS), or gas chromatography mass spectrometry (GC–MS)
- report Limit of Detection and Limit of Quantitation, and measurement uncertainty for all analytical determinations
- provide removal or degradation data demonstrating the effectiveness of processing steps (for example, washing, filtration, heat treatments, or digestion simulations)
- provide data on the impact of cooking or metabolic processing, to demonstrate the extent of inactivation of biologically active components, such as growth factors
3.4.9 Cell line stability
You should show that growth media components and key process parameters do not introduce cell line instability that could make the novel food unsafe.
See our identity, microbiology and production guidance for more information on the impact of growth media components on cell line stability.
4. Assessing toxicology
The toxicological assessment of cell-cultivated products is a core requirement of novel food evaluation. You should address it your application, unless you can scientifically justify an alternative approach.
Toxicological assessments should be based on knowledge of all relevant individual components in the context of the final novel food.
You should justify that the test material is representative of the product as intended to be consumed.
Your should demonstrate that consumption under the proposed conditions of use is not expected to pose a risk to human health. This should consider:
- the properties of the cell biomass
- any novel or modified constituents
- residues of growth media components
- process‑related impurities, metabolites
- substances introduced during manufacturing
Your toxicological evidence should be proportionate to the nature and complexity of the cell-cultivated product, to support an overall conclusion on safety.
4.1 Absorption, distribution, metabolism and excretion (ADME)
You should provide information on the ADME of the toxicologically relevant hazardous components present in the final novel food, to support the toxicological assessment.
You may use other approaches, but these will need justification in your application.
This guidance focuses on ADME considerations for individual components of toxicological relevance.
You should address ADME information related to the nutritional assessment of the final novel food in line with the FSA and FSS guidance on cell-cultivated product allergenicity and nutrition.
Where ADME information is not considered necessary (for example, for substances with negligible bioavailability), you should provide clear scientific justification supported by published literature where available.
4.1.1 Using a tiered approach to ADME assessment
We recommend a tiered, risk-based approach to determine how much ADME information you need for each component used to produce your cell-cultivated product.
This makes sure the assessment is proportionate to the novelty, exposure potential and toxicological relevance of each constituent.
You should clearly justify the selected tier, demonstrate how uncertainties are progressively reduced, and provide supporting scientific evidence at each stage.
You should:
- justify the selection of the tier used
- demonstrate the scientific basis for the chosen approach
- explain how each tier’s findings reduce uncertainty regarding systemic exposure and metabolite formation
You should integrate your ADME tier rationale with the overall toxicology and exposure assessment to support the safety conclusion for the cell-cultivated product.
Tier 1: Existing knowledge and low concern materials
Tier 1 may be sufficient where robust evidence from published literature is available to describe the expected fate of the toxicologically relevant components and their potential metabolites.
You may rely on established digestion pathways, known breakdown of conventional animal cells, and evidence indicating negligible systemic exposure.
This tier is appropriate for components that are well characterised, have a recognised history of safe dietary use (for example, conventional food ingredients), or are present at levels too low to contribute meaningfully to systemic uptake.
No new ADME testing is required at this tier, provided the justification is well supported.
Tier 2: In vitro and in silico assessment
You should progress to Tier 2 assessments if Tier 1 information is insufficient to fully characterise ADME profile (for example, in substances that occur naturally in conventional foods).
These may include in vitro gastrointestinal digestion models, cellular uptake or transport assays, metabolism studies in biorelevant fluids, and in silico modelling approaches to predict absorption, distribution, metabolic pathways, or excretion profiles.
Tier 2 investigations help determine whether any component may become systemically available, produce metabolites of interest, or warrant further evaluation.
Outcomes from this tier should be used to address residual uncertainties and determine whether higher tier testing is required.
Tier 3: Targeted in vivo ADME studies
Tier 3 applies only when findings from Tier 1 and Tier 2 indicate potential systemic exposure formation of metabolites requiring evaluation, or unresolved uncertainty regarding constituent fate.
These may be substances with no history of safe use, but where toxicological data exists to support safety.
You may conduct targeted in vivo toxicokinetic studies designed to address specific data gaps, (for example, tissue distribution, biotransformation pathways and elimination rates).
Studies conducted at this tier should be scientifically justified, directly address remaining uncertainties, and use test materials representative of the marketed cell-cultivated product.
Tier 4: Hazard‑driven, higher‑tier ADME investigations
Tier 4 is reserved for scenarios where specific hazards or safety concerns have been identified, such as:
- the formation of bioactive or potentially toxic metabolites
- evidence of persistence or accumulation
- properties suggesting systemic exposure at levels of concern
This may include substances with no history of safe use and no supportive toxicological data.
Studies at this tier are hazard‑driven and may include advanced in vivo investigations or comprehensive metabolite profiling to fully characterise exposure, metabolic fate and biological relevance.
4.2 Toxicological information
You should assess the toxicological safety of the cell-cultivated product novel food using a structured, weight of evidence approach.
The toxicological strategy may be informed by:
- source, production process, identity and composition of the cell-cultivated product
- existing toxicological evidence, including in silico, in vitro and in vivo studies, human data, and relevant information from other sectors where applicable
- a comprehensive literature review covering the cell-cultivated product constituents and relevant metabolites
- ADME information, including potential systemic exposure
Where data gaps remain after reviewing available evidence, you should conduct appropriate toxicological studies on toxicologically relevant components representative of the cell-cultivated product novel food.
4.2.1 Genotoxicity
You should assess the genotoxic potential of the cell-cultivated product using a stepwise testing strategy consistent with internationally recognised approaches for novel foods. You may also refer to the UK guidance on a strategy for genotoxicity testing of chemicals, to inform your approach to genotoxicity testing.
The following sequence of tests is required unless scientifically justified otherwise:
-
Assessment of existing genotoxicity evidence: this may include in silico, in vitro and in vivo studies, human data, and relevant information from other sectors where applicable.
-
In vitro testing: you should conduct both a bacterial reverse mutation test (Ames test) and an in vitro mammalian cell assay using lysed cell-cultivated product biomass representative of the novel food and components of toxicological relevance.
-
In vivo follow-up: where in vitro results are positive, equivocal or inconclusive, applicants should perform an appropriate in vivo genotoxicity study on toxicologically relevant components. You should integrate genotoxicity findings into the broader toxicological assessment and provide clear justification on how the data supports the safety of the cell-cultivated product novel food as intended to be consumed.
4.2.2 Repeated dose toxicity, including 90-day studies
You should assess the need for in vivo repeated dose toxicity studies on a case-by-case basis. Consider:
- the composition of the cell-cultivated product
- its constituent components
- expected product/component intake and systemic exposure levels
- the completeness of the available safety dataset
Repeated dose studies are required where components lack sufficient toxicological information and are present in the novel food.
Animal models should only be used when a component presents a toxicological concern and necessary safety data cannot be generated using alternative methodologies. Where used, testing should only be conducted on the component of concern.
Where studies are required, you should:
- conduct sub chronic (90-day) oral toxicity studies according to OECD 408, using individual toxicologically relevant components or test material representative of the cell-cultivated product novel food
- consider including a recovery period after the 90 days, as it strengthens the study design
- consider the need for higher tier studies, where applicable (for example, reproductive or developmental toxicity studies or long-term toxicity studies)
4.2.3 Special considerations for proteins (for example, recombinant growth factors)
Where the cell-cultivated product contains proteins or molecules with biological activity, such as recombinant growth factors, cytokines or signalling peptides, you should provide a tailored assessment.
Your assessment should address the following:
- the potential for systemic exposure, including evidence that the protein is denatured, degraded or otherwise rendered inactive under gastrointestinal conditions
- the effect of processing, such as heat, pH changes or mechanical disruption, on protein structure and activity
- evidence demonstrating lack of oral bioavailability, supported by literature, in vitro digestion studies or relevant degradation data
4.2.4 Threshold of Toxicological Concern (TTC)
You may apply the TTC approach to assess substances present in cell-cultivated products at low concentrations where conventional toxicity data are limited. The TTC approach is not appropriate for substances where UK food or feed legislation mandates the provision of toxicological data.
If adequate substance-specific data exist or can be produced to support a full risk assessment, those data should be used in preference to the TTC approach.
You may refer to EFSA guidance on the use of the Threshold of Toxicological Concern approach in food safety assessment.
You may support toxicological assessment using read-across approaches, when using reviewed guidelines by the Toxicological Committee.
4.2.5 Considering vulnerable populations and margin of safety
The toxicological assessment is usually performed based on product intakes in the healthy adult consumer population.
You should consider whether the toxicological assessment is applicable to vulnerable subpopulations, including but not limited to:
- infants, toddlers, children and adolescents
- pregnant or breastfeeding women
- immunocompromised individuals
Where relevant, you should:
- provide justification that toxicological conclusions apply to these groups, considering exposure based on body weight
- apply additional uncertainty factors in the risk assessment, or provide subgroup-specific analysis if toxicological endpoints suggest heightened sensitivity
- ensure that margins of safety remain protective across all intended consumer groups
4.2.6 Data from studies involving humans
You may provide any human data relevant to the safety assessment of the cell-cultivated product, where available.
Human studies are not generally considered safety studies in their own right. They are typically conducted as food tolerability studies once an acceptable safety case has been established.
Where available, human data can form an important component of the overall weight of evidence, providing valuable information on the tolerability and consumption of a novel food under anticipated conditions of use.
Human intervention studies should be included whenever biological measures are assessed that could be relevant to the interpretation of safety evidence, even if the primary purpose of the study was not safety (for example, efficacy trials or taste trials).
Observational human data, where available, may also provide supportive product tolerability information and should be summarised accordingly.
You should present data according to a hierarchy of evidence, reflecting study quality and relevance. You should integrate human findings with the wider toxicological dataset to support the overall safety conclusion for the cell-cultivated product novel food.
4.2.7 Weight of evidence and overall toxicological conclusions
You should integrate all available evidence to form an overall conclusion on the safety of the cell-cultivated product.
To form an overall conclusion, include:
- literature evidence
- genotoxicity
- repeated dose studies
- use of TTC assessments
- ADME profile
- exposure estimates
The weight of evidence narrative should:
- clearly articulate how potential hazards have been addressed
- identify any remaining uncertainties and explain why they do not impact safety under the proposed uses
- propose specification limits for constituents or impurities requiring control, such as residual growth factors or components with biological activity, antimicrobial residues and heavy metals
Figure 1: Flowchart showing a toxicological assessment process.
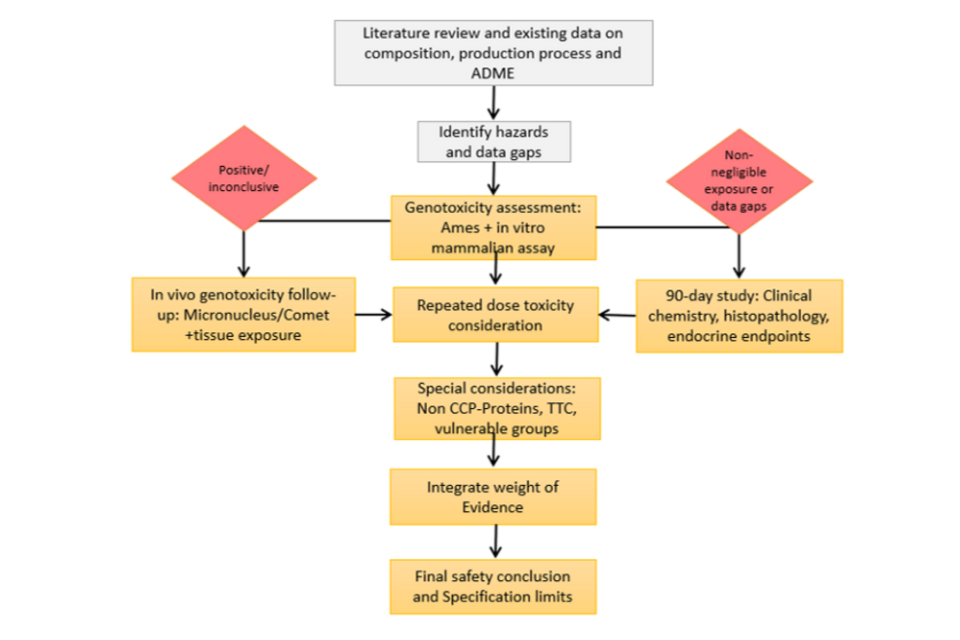
Figure 1 flowchart alternative text: The process begins with a literature review and existing data on composition, production process and ADME. This information is used to identify hazards and data gaps, followed by a genotoxicity assessment using the Ames test and an in vitro mammalian assay.
If results are positive or inconclusive, an in vivo genotoxicity follow-up is carried out, including micronucleus and tissue exposure studies, before returning to repeated dose toxicity consideration.
If there is non-negligible exposure or significant data gaps, a 90-day study is conducted examining clinical chemistry, histopathology and endocrine endpoints, and the findings are also fed into repeated dose toxicity consideration.
The process then moves to assessment of special considerations, including non-CCP proteins, the Threshold of Toxicological Concern (TTC), and vulnerable groups. All evidence is integrated using a weight-of-evidence approach, leading to a final safety conclusion and specification limits.
5. Concluding your application
You should bring together the information requested across all sections into a concise overall assessment. This should explain how it supports the safety of the cell-cultivated product under the proposed conditions of use.
6. Acknowledgements
Members of the Advisory Committee on Novel Foods and Processes (ACNFP) and the Subcommittee on Cell Cultivated Products who provided expert views at the cell-cultivated product subgroup meeting CCP03/05 (September 2025 and April 2026) and reviewed this guidance as part of the ACNFP June 2026 meeting.